Mediterranean anemia is also known as thalassemia. Its types include alpha and beta thalassemia. This condition occurs as a result of gene mutations. These mutations occur in genes affecting the structure of the hemoglobin molecule and affect hemoglobin production.
Symptoms usually begin to appear in childhood or early adulthood. Symptoms include fatigue, pale skin, weakness, shortness of breath, and rapid heartbeat. Mediterranean anemia can significantly affect the quality of life of affected individuals and requires a long-term treatment plan and follow-up checks.
What is Mediterranean Anemia (Thalassemia)?
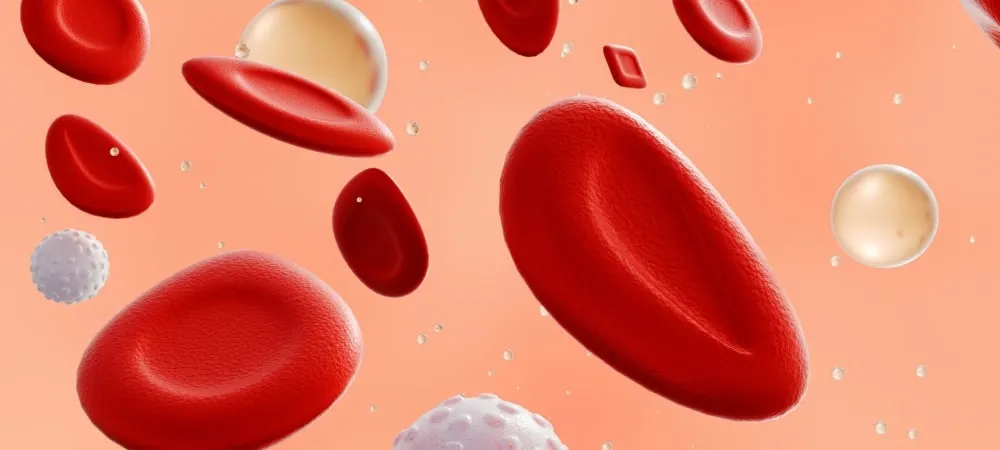
Mediterranean anemia is a condition that occurs due to abnormalities in the production of hemoglobin found in the blood. Hemoglobin enables red blood cells to carry oxygen throughout the body. Due to problems encountered in the transport of oxygen, anemia may occur in individuals, which can lead to symptoms such as weakness, pallor, fatigue, shortness of breath, bone pain, and growth and development retardation.
What Are the Symptoms of Mediterranean Anemia?
Symptoms may vary depending on the type of thalassemia seen in individuals. In cases where a single alpha gene is missing, symptoms may often not be observed. Some of the symptoms of thalassemia may include:
Dizziness
Headache
Shortness of breath
Rapid heartbeat
Fatigue
Weakness
Growth problems
Delayed puberty
Bone abnormalities
Enlargement of the spleen
Pale skin
Yellowish skin
Loss of appetite
Irregular bone structure in the face
Change in urine color
What Causes Mediterranean Anemia?
Thalassemia is fundamentally a genetic disorder. This condition occurs when a person inherits mutated genes that alter hemoglobin from their parents. Mutations in the genes in the alpha or beta globin chains that make up the hemoglobin molecule cause the disorder. These mutations affect normal hemoglobin production in the body and lead to anemia. If both of your parents have thalassemia, you may have it too. If you inherit two or more abnormal gene copies from your parents, mild to severe symptoms may occur depending on which protein is affected.
What Are the Types of Mediterranean Anemia (Thalassemia)?
There are two types of thalassemia disorders: alpha thalassemia and beta thalassemia. Each type of thalassemia may exhibit different symptoms and severity levels. Diagnosis and treatment approaches vary depending on the type and severity of the disease. Types of thalassemia may include:
Alpha Thalassemia
Alpha thalassemia is caused by mutations in the alpha globin chains. This is generally common in Southeast Asia, the Middle East, and Africa. Four genes are inherited, two from each parent, which make up the alpha globin protein chains. This type of anemia develops when there is one or more abnormal genes. The number of defective genes inherited determines whether symptoms of anemia will occur and, if so, how severe those symptoms will be.
Beta Thalassemia
Beta thalassemia occurs due to mutations in the hemoglobin beta chains. Individuals receive one beta-globin gene from each of their parents. This disorder is seen in two main forms: beta thalassemia major and beta thalassemia minor. Beta thalassemia major is the most severe form and usually shows symptoms in the first few years of life. Beta thalassemia minor generally presents with milder symptoms.
How is Mediterranean Anemia Diagnosed?
In thalassemia, blood values obtained from patients, in particular, can provide useful information for diagnosis. Doctors may request various tests to diagnose the disorder. Some of these tests are:
Complete blood count (CBC): Individuals with thalassemia have fewer blood cells and lower hemoglobin levels than the healthy red blood cell and hemoglobin values found in normal individuals. In addition, the sizes of red blood cells are smaller compared to healthy red blood cells.
Reticulocyte count: This test can show whether there are fewer red blood cells in the bone marrow than should be in the body.
Iron measurement tests: This can be a useful test in determining the cause of anemia. It can show whether the anemia seen in individuals is iron deficiency or thalassemia.
Hemoglobin electrophoresis: This test is used to identify different types of hemoglobin. In individuals with this disorder, hemoglobin variants different from normal may be detected.
Genetic tests: Genetic tests can be performed to confirm the diagnosis. These tests are used to identify specific gene mutations for which the patient is a carrier or affected.
Additionally, in rare cases, a bone marrow biopsy may be requested by the doctor to confirm the diagnosis. In this procedure, a bone marrow sample is taken and examined. In individuals with thalassemia, structural abnormalities of red blood cells in the bone marrow can be detected.
How is Mediterranean Anemia Treated?
Mild forms of thalassemia generally do not require treatment. Treatments for moderate to severe thalassemia usually involve blood transfusions and iron chelation therapy. Mediterranean anemia treatment may include:
Blood transfusion: Individuals with thalassemia often need regular blood transfusions. Blood transfusions help alleviate anemia symptoms and improve quality of life. In blood transfusions, the method of intravenous injection of red blood cells is applied to bring healthy red blood cells and hemoglobin to normal levels.
Iron chelation therapy: Continuous blood transfusions can cause excessive iron accumulation in the body. Too much iron can damage organs. Therefore, it is important to remove excess iron from the body using methods such as iron chelation therapy. This treatment is usually carried out using medications or iron binders.
Folic acid supplementation: Folic acid is important for the production of red blood cells and plays an important role in the body's production of healthy blood cells. People with Mediterranean anemia often take folic acid supplements because low folic acid levels can worsen anemia symptoms.
Bone marrow and stem cell transplantation: Bone marrow transplantation and stem cell transplantation may be recommended for individuals with severe Mediterranean anemia. This procedure aims to promote normal production of red blood cells by transplanting healthy bone marrow cells. The doctor applies the treatment by injecting bone marrow stem cells taken from the donor into the patient during the procedure.
Frequently Asked Questions?
Mediterranean Anemia is generally handled by Hematology or Pediatric Hematology specialists. These specialists are involved in the diagnosis and treatment of blood diseases. The Mediterranean anemia blood test generally comes out within 2-3 days. However, depending on the workload of the laboratory where the test is performed, the test methods used, and the type of test, results may take a different amount of time. A DNA test can take about 4-5 days. Mediterranean anemia is a genetic disease. It passes from parents to child in an autosomal recessive manner. It occurs due to specific mutations in the beta globin gene. These mutations lead to abnormalities in hemoglobin production and consequently to anemia. It is transmitted genetically. Mediterranean Anemia is not a type of cancer. Mediterranean Anemia is a condition that occurs due to abnormalities in the production of red blood cells. Cancer, on the other hand, is the uncontrolled growth and spread of cells in different tissues of the body. Mediterranean Anemia treatment aims to alleviate the symptoms of anemia and increase the quality of life, whereas cancer treatment aims to destroy or control cancer cells. Both conditions are different health problems with different pathophysiology. Although Mediterranean Anemia and anemia (low blood count) have similar symptoms, they are actually disorders indicating different conditions. Mediterranean Anemia is a type of disorder that causes red blood cells to be produced in fewer numbers than normal or in an ineffective manner. Anemia, on the other hand, is the condition of decreased oxygen-carrying capacity in the body as a result of a decrease in the number or function of red blood cells (hemoglobin) in the blood, and this condition can arise for many different reasons. If the disorder is not treated, disease symptoms may worsen over time and negatively affect the quality of life. Iron accumulation may occur in the body due to chronic blood transfusions, which can lead to organ damage, especially liver and heart problems. Mediterranean Anemia carrier state refers to the condition where a person genetically carries the abnormal hemoglobin gene but does not show symptoms. In this case, individuals have specific mutations in the beta globin gene, but symptoms of the disease are generally absent. Carriers may be healthy, but they can potentially pass on thalassemia disease due to the mutations they carry. Beta thalassemia is a type of blood disease that leads to the production of red blood cells in fewer numbers than normal or in an ineffective manner due to a genetic disorder in the hemoglobin beta chains. The definitive treatment for thalassemia is a bone marrow transplant from a compatible family member. Problems finding a suitable donor and serious complications of the transplant procedure are risk factors encountered during the treatment process. However, treatments aimed at controlling symptoms can be planned. Treatment options include regular blood transfusions and medications to prevent iron deficiency.
Mediterranean Anemia (Thalassemia) is a genetic blood disease that can be controlled with early diagnosis and proper follow-up. If you or your child is suspected of having thalassemia, regular blood checks and evaluation by a specialist physician are of great importance.
You can apply to our Internal Medicine outpatient clinic to get detailed information and make an appointment.
Start with the right assessment for What is Mediterranean Anemia (Thalassemia)? Symptoms, Causes and Treatment
Share your symptoms with our specialist physicians; let us determine the right diagnosis and treatment plan together.
Warning
If your symptoms started suddenly, are progressing rapidly, or significantly restrict your daily life, consult a physician without delay.
Plan your What is Mediterranean Anemia (Thalassemia)? Symptoms, Causes and Treatment journey with our specialists.
Fill in the form or book an appointment to learn about treatment options, the process and preparation.